


Every GMP site relies on Qualification and Validation to protect product quality and ensure compliance. Even a mid-size facility manages many qualified systems and validated processes, so a clear understanding of Qualification vs Validation in Pharma is essential to avoid delays and deviations. For deeper strategy and real project flows, you can explore the Pharma Validation page on Pharmuni.
Qualification confirms that equipment and facilities work as intended, while validation proves that processes deliver consistent results. This guide explains four key differences between Qualification and Validation in Pharma using simple language, real industrial examples, and practical GMP documentation insights.
Table of Contents
What Is Qualification in Pharma?
Qualification is the structured process used to prove that facilities, utilities, and equipment can operate safely and reliably under routine GMP conditions. It applies primarily to physical systems that support manufacturing and quality control.
Typical systems covered by qualification include:
- Manufacturing equipment (mixers, granulators, autoclaves)
- HVAC and environmental control
- Utilities like purified water, steam, compressed air, and gases
- Supporting infrastructure such as cold rooms and warehouses
What Is Validation in Pharma?
Now move from “tool” to “recipe”. Validation confirms that a process, method, or cleaning procedure delivers consistent, compliant output over time. Regulators describe process validation as a lifecycle with design, qualification, and continued verification.
Typical validation areas include:
- Process Validation for manufacturing steps
- Cleaning Validation for shared equipment
- Analytical Method Validation in QC labs
- Computer System Validation (CSV) for GxP software and data
Is Qualification Part of Validation? What the Guidelines Say
Yes. Qualification is a building block of validation, but it is not the same thing. The ICH Q7 GMP guide for active pharmaceutical ingredients defines qualification as the action of proving and documenting that equipment or ancillary systems are properly installed, work correctly and actually lead to the expected results. It then adds that qualification is part of validation, but the individual qualification steps alone do not constitute process validation.
EU GMP Annex 15 makes the same link in practical terms. It describes the qualification stages for equipment, facilities, utilities and systems, from design qualification through IQ, OQ and PQ. In its process validation section, it states that equipment, facilities, utilities and systems used for process validation should be qualified. In other words, a qualified autoclave is a precondition for a validated sterilisation process, not proof of it.
This distinction matters when you prepare for an inspection. A complete set of qualification reports shows that your tools work as intended. However, it does not show that your process consistently delivers product that meets its specifications. That second claim needs process validation data.
Key Differences Between Qualification and Validation
At first glance, both processes look similar: protocols, testing, reports, and QA approvals. However, Qualification vs Validation in Pharma differs in several important ways:
- Scope Difference
- Output Difference
- Documentation Difference
- Risk & Compliance Impact
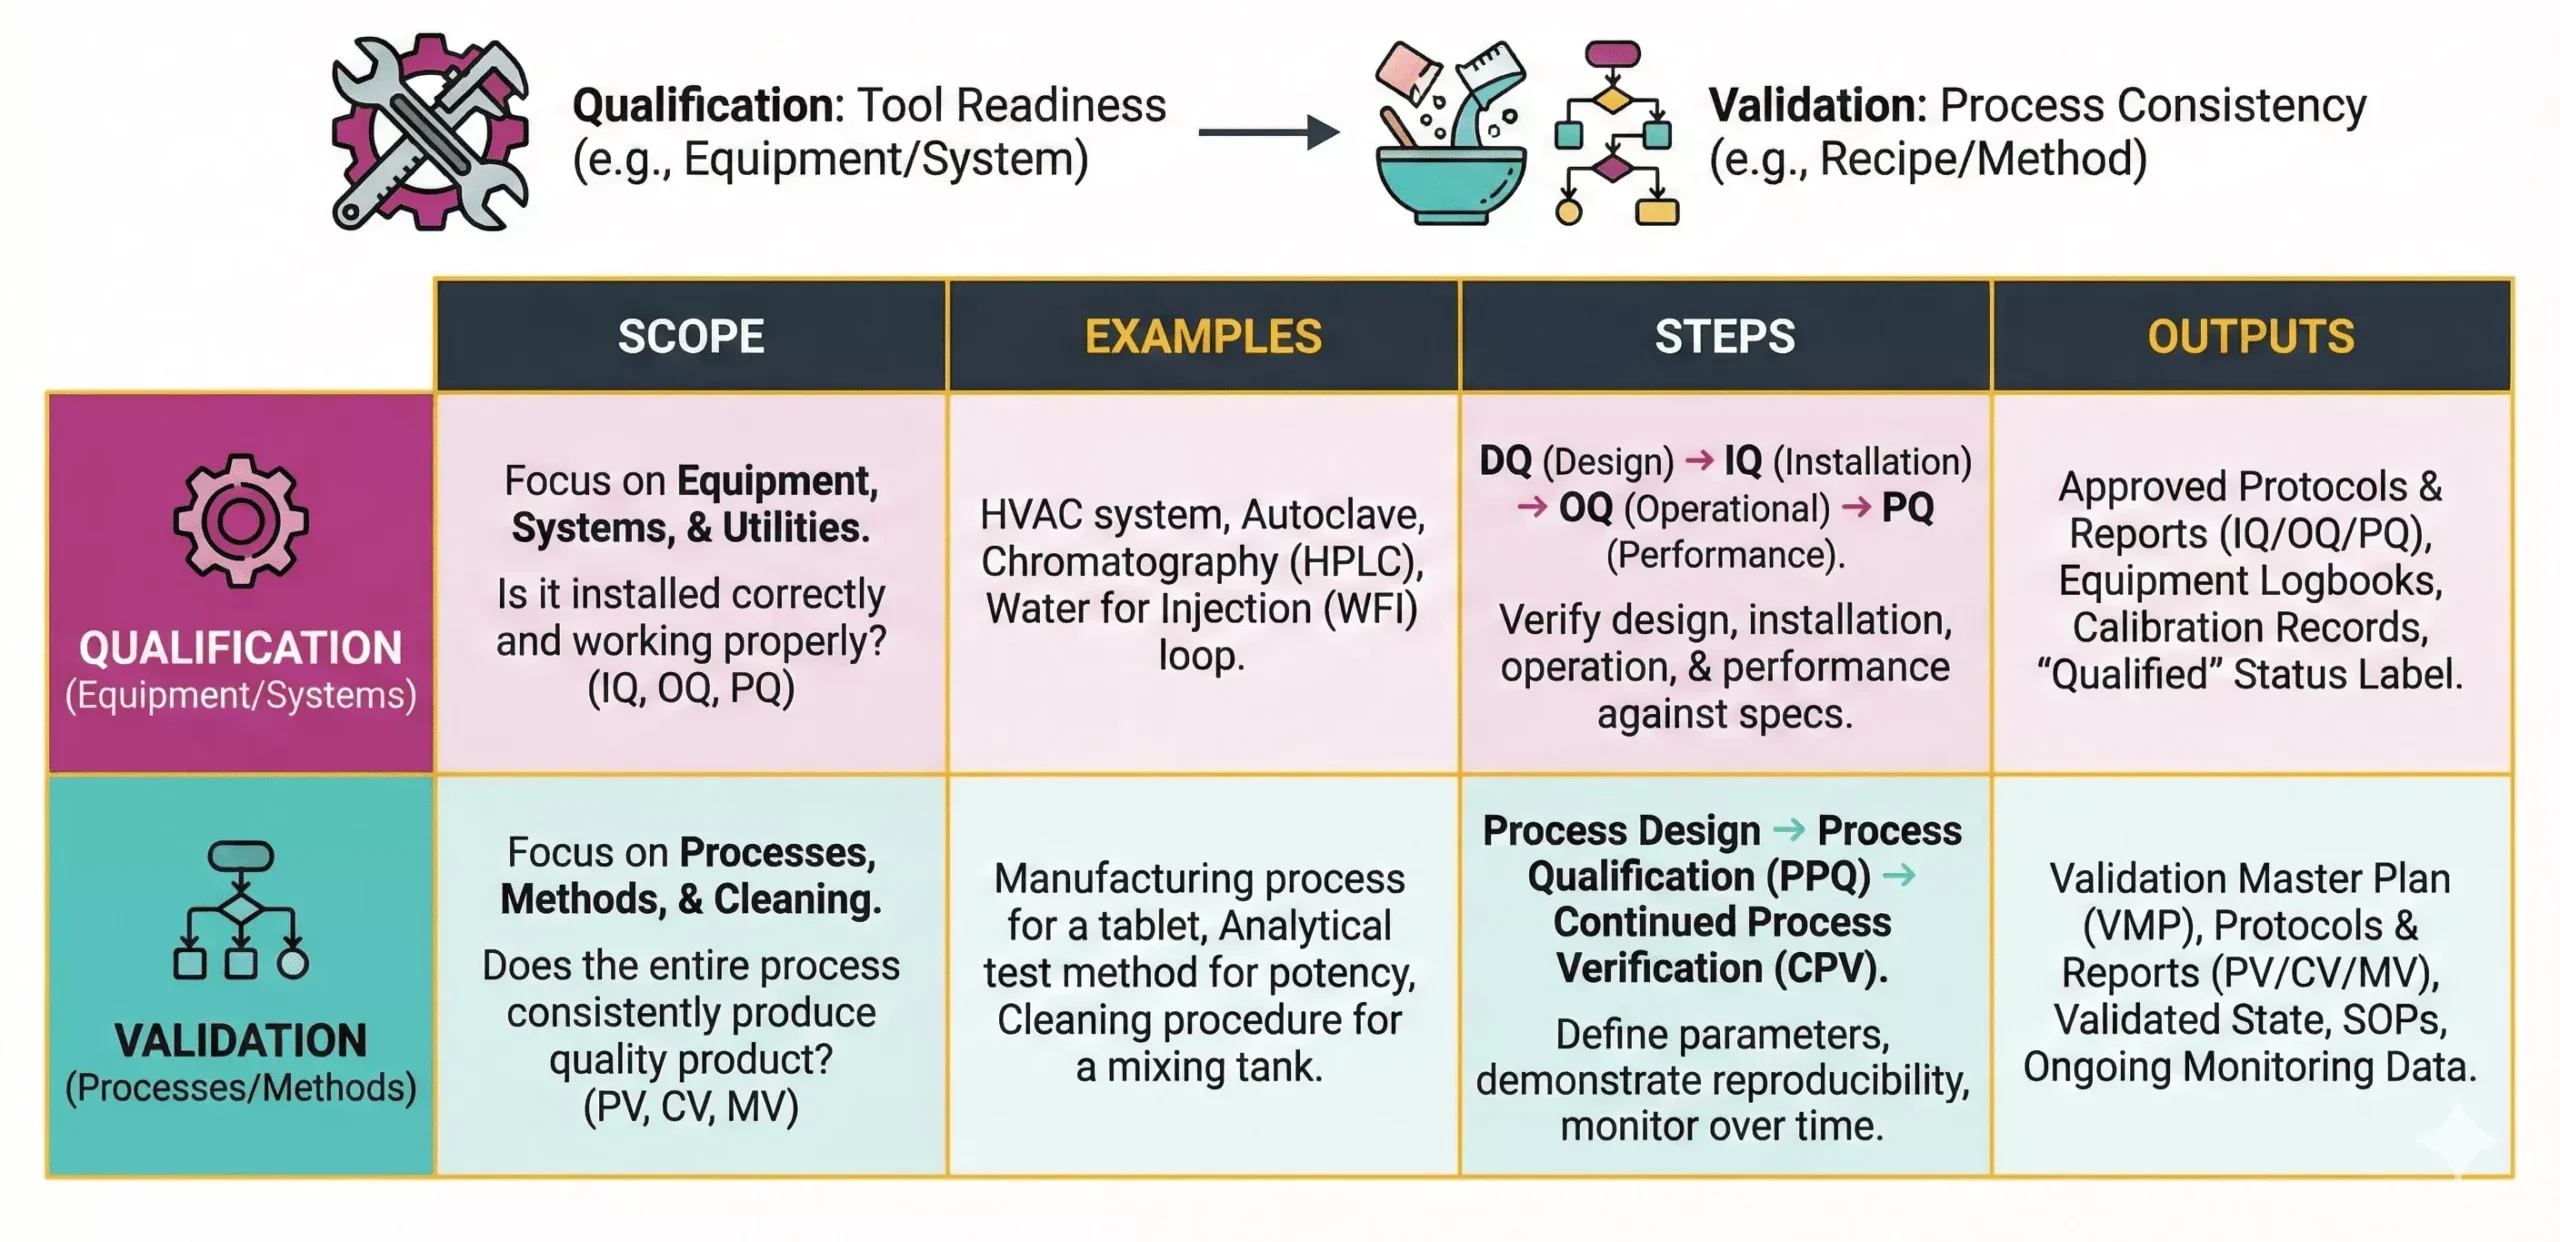
1. Scope Difference
Qualification
- Qualification focuses on equipment, utilities, facilities, and systems.
Validation
- Validation focuses on processes, methods, cleaning, and software use.
- Equipment, facilities, utilities and systems used for process validation should be qualified first (EU GMP Annex 15, section 5.9).
2. Output Difference
Qualification output:
- Evidence that systems meet user requirements
- Defined operating ranges and limits
- A documented qualified state for GMP use
Validation output:
- Proof that processes consistently meet specifications
- Control of CPPs and CQAs
- Trending data through CPV to confirm ongoing consistency
3. Documentation Difference
Equipment Qualification Documentation
- URS and functional specifications
- DQ, IQ, OQ, and PQ protocols and reports
- FAT/SAT records
- Calibration and maintenance logs
Validation documentation:
- Validation Master Plan (VMP)
- Process validation and PPQ protocols
- Cleaning validation plans and matrices
- CPV reports and risk assessments
4. Risk & Compliance Impact
Qualification Risk Impact:
- Poor qualification leads to equipment failures, downtime, and data integrity issues.
Validation risk impact:
- Poor validation leads to batch failures, recalls, and patient safety risks.
Both activities are essential to control GMP risk and withstand regulatory inspections.
Qualification and Validation Lifecycles
The lifecycle view helps teams avoid “one-and-done” thinking. Both qualification and validation run as ongoing cycles.
- Equipment Lifecycle
- Process Validation
- When Requalification Is Needed
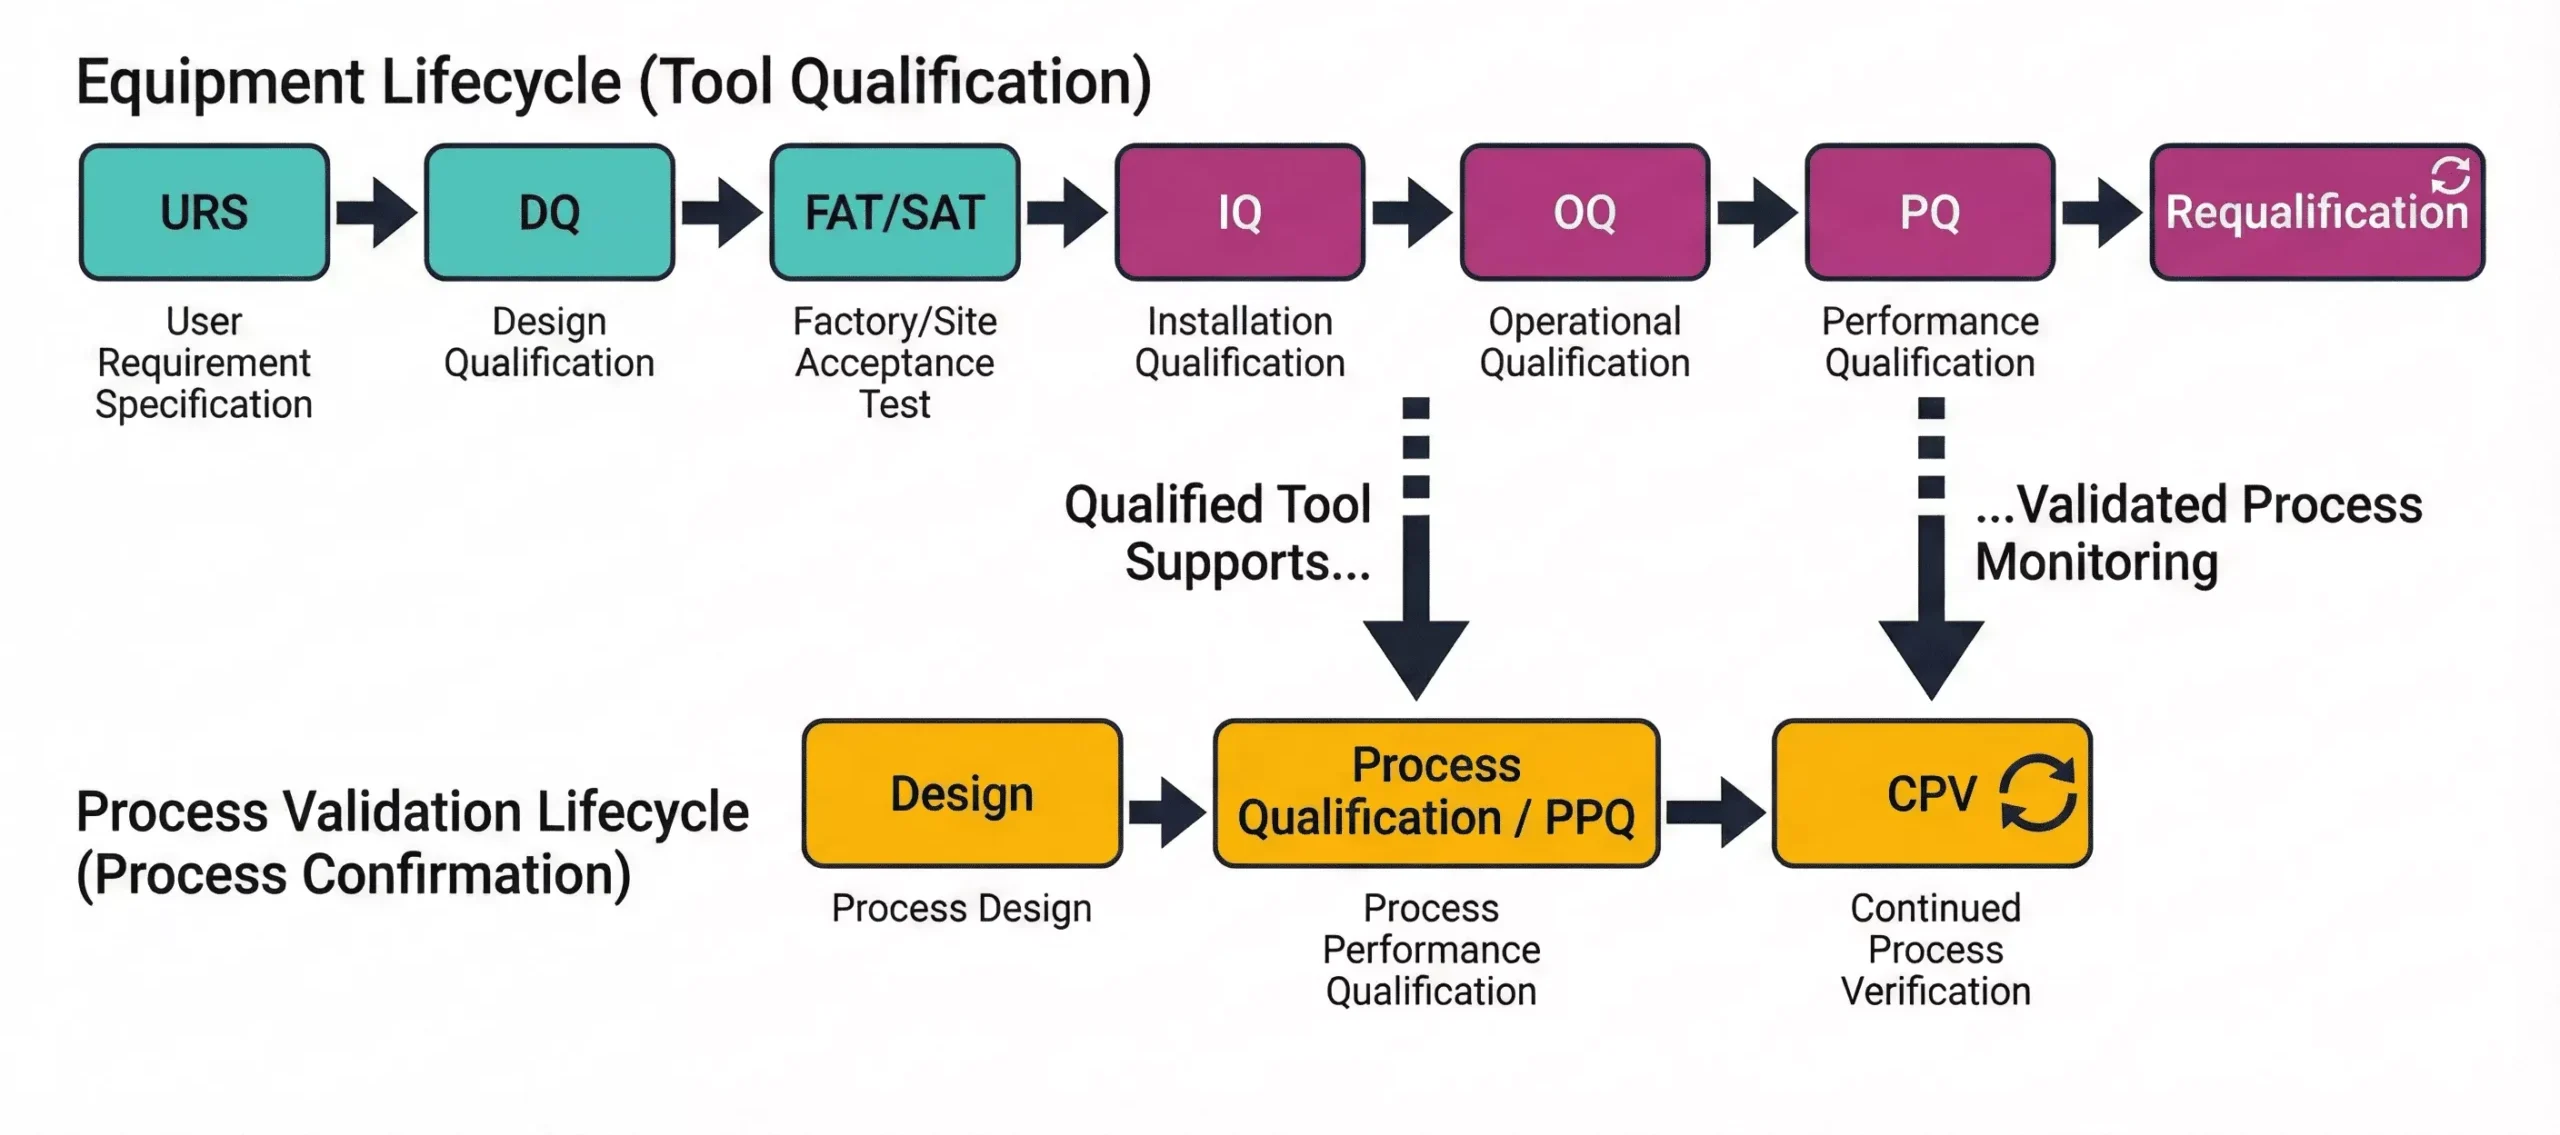
Equipment Lifecycle
A typical Equipment lifecycle includes:
- Concept and URS
- Vendor selection, FAT / SAT
- DQ, IQ, OQ, PQ
- Routine operation with maintenance and calibration
- Periodic review and requalification
- Decommissioning and retirement
Process Validation Lifecycle
A typical Process Validation lifecycle includes:
- Process development and scale-up
- Risk assessment and control strategy design
- Process qualification / PPQ batches
- Continued process verification with trending
- Ongoing improvement and change control
When Requalification Is Needed
Qualification does not end with PQ. Annex 15 expects equipment, facilities, utilities and systems to be evaluated at an appropriate frequency to confirm that they remain in a state of control. If a site requalifies on a fixed schedule, it should justify the interval, define the evaluation criteria and assess whether small changes have built up over time. Utilities need extra care, because the period and extent of their qualification should reflect seasonal variations where relevant. In addition, every planned change should pass through change control, where quality risk management decides whether new validation, verification or requalification work is required.
Where Qualification Sits in the FDA Validation Lifecycle
The FDA’s 2011 process validation guidance shows exactly where qualification fits. It defines process validation as the collection and evaluation of data, from the process design stage through commercial production, which establishes scientific evidence that a process is capable of consistently delivering quality products. The guidance then organises this work into three stages.
Stage 1 – Process Design: The commercial manufacturing process is defined based on knowledge gained through development and scale-up.
Stage 2 – Process Qualification: The process design is evaluated to confirm it can deliver reproducible commercial manufacturing. This stage covers the design of the facility and the qualification of utilities and equipment, followed by process performance qualification (PPQ).
Stage 3 – Continued Process Verification: Ongoing assurance is gained during routine production that the process remains in a state of control.
Equipment and utility qualification therefore sits inside Stage 2, before the PPQ batches. EU GMP Annex 15 calls the equivalent of Stage 3 ongoing process verification and describes it as documented evidence that the process remains in a state of control during commercial manufacture. For a stage-by-stage walkthrough, see our guide to process validation in pharmaceutical.
Real Industrial Examples
Real stories make these concepts stick. Here are three classic scenarios that show how Qualification and Validation in Pharma works on the shop floor.
Example 1: Equipment Qualification – Autoclave for Terminal Sterilization
A sterile manufacturing site installs a new autoclave. The team:
- Defines URS for cycles, loads, temperatures, and data recording
- Performs FAT to test controls and safety features
- Executes IQ to verify installation and calibration
- Runs OQ to map worst-case conditions
- Conducts PQ using real or simulated loads
Only after qualification can the sterilization process be validated.
Example 2: Process Validation – Solid Oral Dose Granulation
A tablet plant wants to validate a wet-granulation process. The team:
- Defines CPPs and CQAs from development data
- Executes PPQ batches at commercial scale
- Monitors blend uniformity, moisture, and compression behavior
- Releases product only after successful PPQ results
Here, equipment qualification supports validation, but the process remains the focus.
Example 3: Cleaning Validation and CIP Qualification
On shared equipment:
- Qualification confirms CIP systems, spray balls, and instruments function correctly.
- Cleaning Validation proves residue removal to acceptable limits.
Key activities include swab and rinse sampling, visual inspections, and worst-case product selection.
Cleaning verification is a different activity again. Annex 15 defines pharma cleaning validation as documented evidence that an approved procedure will reproducibly remove the previous product or cleaning agents below the scientifically set maximum allowable carryover. Cleaning verification, by contrast, gathers chemical evidence after each batch or campaign that residues are below that limit. Verification after every batch may be needed while a validation programme is still running, for example for investigational products. However, repeating cleaning and retesting until results pass is not acceptable, and a visual check alone is not enough as an acceptance criterion.
GMP Documentation Required for Both
Documentation forms the backbone of qualification and validation. It ensures traceability, supports audits, and reduces compliance risk.
Both streams usually rely on these shared elements:
- Validation Master Plan (VMP)
- Risk assessments
- SOPs and work instructions
- Training records
- URS and design reviews
- FAT/SAT records
- IQ, OQ, PQ protocols and reports
- Calibration and maintenance records
- Process and cleaning validation protocols
- PPQ documentation
- CPV reports
- Change impact assessments
Strong document linkage supports fast investigations and smooth inspections.
Qualification and Validation: Summary Table
| Aspect | Qualification in Pharma | Validation in Pharma |
|---|---|---|
|
Main focus |
Facilities, utilities, systems, and equipment |
Processes, methods, cleaning, and software |
|
Key question |
“Can this tool operate as intended?” |
“Does this process deliver consistent, compliant product?” |
|
Typical steps |
URS → DQ → FAT/SAT → IQ → OQ → PQ → requalification |
Process design → process qualification / PPQ → continued process verification |
|
Examples |
Autoclave, HVAC, water system, tablet press |
Granulation, sterile filling, cleaning cycles, HPLC method |
|
Primary outputs |
Qualified state, operating ranges, equipment lifecycle control |
Validated state, proven process capability, CPV trends |
|
Main risks |
Downtime, data integrity, technical failures |
Batch failures, recalls, patient safety issues |
|
Key documents |
URS, DQ, IQ/OQ/PQ protocols and reports, FAT/SAT, calibration |
VMP, risk assessments, PV/PPQ protocols, CPV and annual reviews |
|
Regulatory anchor |
EU GMP Annex 15 (sections 3–4); ICH Q7 glossary definition |
FDA Process Validation Guidance (2011); EU GMP Annex 15 (section 5) |
The Annex 15 Revision for Active Substance Manufacturers
Annex 15 itself is changing. Today it applies to medicinal product manufacture and serves only as optional supplementary guidance for active substance (API) manufacturers. In January 2026, the EMA GMP/GDP Inspectors Working Group and PIC/S published a concept paper proposing to make it applicable to manufacturers of chemical and biological active substances. The proposal follows lessons learnt from N-nitrosamine impurities in sartan medicines, and it would also align the annex with ICH Q9(R1) on quality risk management.
Under the proposed timetable, a draft guideline is due for public consultation in December 2026, with publication by the European Commission expected in April 2028. API teams should therefore start mapping their current qualification and validation practices against Annex 15 now.
Common Qualification and Validation Mistakes
Qualification and validation programs often fail because teams treat them as documentation exercises rather than controlled lifecycle activities. One common problem is starting process validation before equipment, utilities, or computerized systems are fully qualified. Weak acceptance criteria also create uncertainty during protocol execution and review. In addition, teams may rely too heavily on vendor documents without confirming site-specific requirements, operating ranges, and GMP risks.
Starting validation before completing required IQ, OQ, or PQ activities.
Using vague, unmeasurable, or unapproved acceptance criteria.
Copying vendor protocols without site-specific risk assessment.
Closing deviations without documenting impact, investigation, and corrective actions.
Failing to reassess qualification or validation after major changes, recurring failures, equipment relocation, software updates, or adverse performance trends.
Final Words
Qualification and validation are essential to maintaining GMP compliance and consistent pharmaceutical product quality. Understanding the different roles of qualification and validation helps teams reduce avoidable risks, prevent project delays, and demonstrate that equipment and manufacturing processes are fit for their intended purposes.
Strong documentation, scientific risk assessment, and lifecycle control support both activities. When qualification and validation are properly planned, executed, and maintained, manufacturers can reduce the likelihood of batch failures and rework while improving inspection readiness.
FAQs
EU GMP Annex 15 says a minimum of three consecutive batches made under routine conditions is generally acceptable for traditional process validation. However, each manufacturer must determine and justify the number based on quality risk management, and a different number can be justified.
No. Annex 15 states that retrospective validation is no longer an acceptable approach. Processes using the traditional approach should be validated prospectively, before the product is certified, and concurrent validation is reserved for exceptional, justified cases.
The qualification process typically follows the DQ–IQ–OQ–PQ sequence:
- Design Qualification (DQ)
- Installation Qualification (IQ)
- Operational Qualification (OQ)
- Performance Qualification (PQ)
Yes. ICH Q7 states that qualification is part of validation, but the individual qualification steps alone do not constitute process validation. Qualified equipment is a prerequisite; process validation must still show that the process consistently delivers quality product.
References
Stephanie Männicke
Digital Marketing Especialist at Zamann Pharma Support, brings 8 years of experience in Corporate and Digital Communication. Specializing in Digital Marketing and Content Creation, Stephanie is currently focused on creating strategic content for Pharmuni's networks, especially content on topics such as recruitment, onboarding and employer branding. Outside of work, Stephanie is a mum, a crocheter and a movie fan. An avid reader and in search of expanding her knowledge, Stephanie is always looking for ways to innovate communication in the digital environment and connect people in a genuine way.
Protocol Deviations in Clinical Trials: Build an Audit-Ready Record
Protocol deviations can threaten participant protection and trial reliability. This practical guide explains risk-based classification, documentation, escalation, CAPA, and preventive oversight.
eCTD Publishing Software: The Right Features for Submission Success
Discover how eCTD publishing software organizes regulatory dossiers, validates electronic submissions, manages document lifecycles, and supports FDA and EMA workflows.
Pharmaceutical Analytical Testing Services: Essential Quality Insights
A practical guide to pharmaceutical analytical testing services: assay, impurities, dissolution, stability, microbial testing, validation and outsourced-laboratory oversight.