


Process Validation in Pharmaceutical means using data across development and manufacture to prove a process consistently delivers quality product. FDA’s first guideline in 1987 and its lifecycle-based update in 2011 define three stages: process design, process qualification, and continued process verification, all embedded in cGMP expectations.
Pharma Validation matters because regulators link it directly to patient safety and product reliability.
EU GMP Annex 15, revised in 2015 and used together with the EMA guideline on process validation for regulatory submissions, aligns with FDA expectations by requiring risk-based, lifecycle validation. This framework guides modern GMP worldwide.
Table of Contents
What Is Process Validation in Pharmaceuticals?
Process validation in pharmaceuticals means you control how a process runs from start. You link science, risk management, and solid documentation.
FDA defines process validation as the collection and evaluation of data, from the process design stage through commercial production, which establishes scientific evidence that a process is capable of consistently delivering quality product. EU GMP Annex 15 describes it as documented evidence that the process, operated within established parameters, can perform effectively and reproducibly to produce a medicinal product meeting its predetermined specifications and quality attributes.
Process validation stages include process design, process qualification, and continued process verification. You focus on critical quality attributes and process parameters. Moreover, you align process validation with GMP rules and inspection expectations worldwide. Regulators expect strong pharmaceutical validation to lower risks and protect thousands of future patients. You might reduce major deviations from 4 each year to 1 after improvements. You also compare 100% critical parameter monitoring with earlier sampling checks on 10% data.
Regulators expect strong pharmaceutical validation to lower risks and protect thousands of future patients. You might reduce major deviations from 4 each year to 1 after improvements. You also compare 100% critical parameter monitoring with earlier sampling checks on 10% data.
Regulatory Framework for Process Validation
Regulatory frameworks guide process validation from early design through commercial manufacturing. Authorities link validation with GMP rules to protect patients and ensure reliable quality.
Moreover, FDA guidance and EMA Annex 15 describe lifecycle expectations for process validation stages. You design, qualify, and monitor processes using science-based, risk-based principles.
Therefore, companies align internal procedures with these frameworks to meet global inspection expectations.
- FDA Guidelines (FDA 2011 Process Validation Guidance)
- EMA/EU Guidelines (EU Annex 15 Overview)
- WHO & ICH References
FDA Guidelines (FDA 2011 Process Validation Guidance)
FDA process validation guidance from 2011 explains how manufacturers design reliable processes. It links process understanding and control strategies to protect patients.
Therefore, companies follow FDA process validation stages and adjust processes when data change. Teams share findings across sites.
EMA/EU Guidelines (EU Annex 15 Overview)
EU Annex Fifteen explains how to plan process validation activities. Moreover, it links qualification steps with GMP requirements for facilities and equipment.
EMA expects firms to apply risk management when they design validation. The guideline from 2015 supports lifecycle thinking and consistent product quality.
Key points from EU Annex 15:
Define clear process validation stages and responsibilities.
Link qualification, cleaning, and transport validation.
Use documented risk assessments for critical steps.
Keep evidence ready for EMA and local inspectors.
WHO & ICH References
WHO guidelines support safe, effective medicines through modern global quality standards worldwide. Moreover, they encourage strong process validation in pharmaceutical manufacturing and distribution.
ICH documents harmonize technical expectations between major regions and regulators. They guide pharma validation strategies and lifecycle approaches for complex products.
WHO references:
WHO GMP guidelines for pharmaceutical products
WHO guidance on quality risk management and inspections
ICH references:
ICH Q8, Q9, Q10 on quality and risk management
ICH Q11 on development and manufacture of drug substances
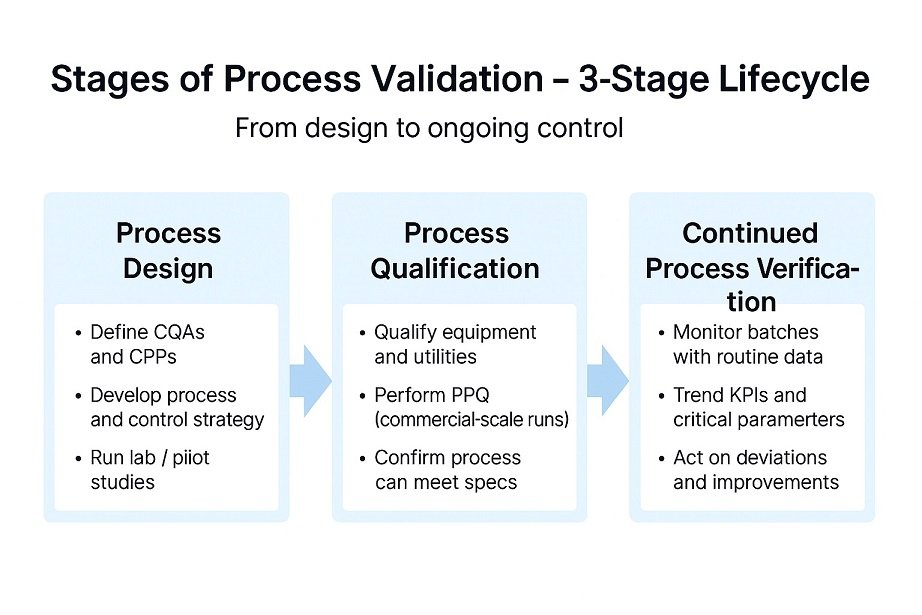
The Three Stages of Process Validation (PV Lifecycle Model)
Process validation stages cover design, qualification, and ongoing verification. You use data to keep every critical step under control.
Moreover, Stage one focuses on process design and key risks. You define parameters and link them directly to quality attributes.
Therefore, Stage two qualifies equipment and utilities for production. Stage three then verifies performance with trends and routine checks.
- Stage 1: Process Design
- Stage 2: Process Performance Qualification (PPQ)
- Stage 3: Continued Process Verification (CPV)
Stage 1: Process Design
Stage 1, Process Design, explains how the process should run. You map inputs and controls across the full PV lifecycle.
Therefore, you define critical parameters and ranges using science reliably. These choices support qualification later and enable continued process verification.
key actions
Define process steps, inputs, and equipment clearly.
Link parameters to quality attributes for later stages.
Stage 2: Process Qualification (Including PPQ)
Stage 2, Process Performance Qualification, runs PPQ batches under routine commercial conditions. Moreover, results confirm that equipment, utilities, and controls work together within defined ranges.
Before PPQ starts, the equipment, facilities, utilities and systems involved must be qualified, and test methods must be validated for their intended use; the differences are explained in our guide to qualification vs validation. PPQ batches should normally match the intended commercial batch size, suppliers of critical starting and packaging materials should be qualified, and production personnel should take part in manufacturing the batches.
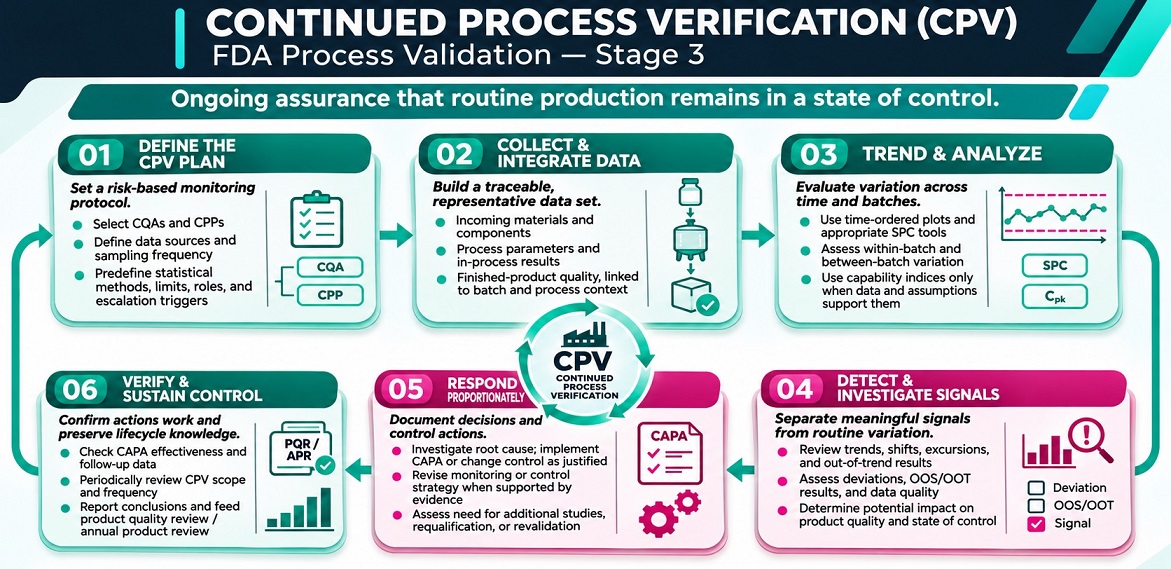
Stage 3: Continued Process Verification (CPV)
Stage 3, Continued Process Verification, monitors performance data over time. Teams track batches and deviations in routine manufacturing.
Moreover, CPV identifies trends early and supports process adjustments. Strong monitoring maintains control and protects patient safety.
Digital Tools in Process Validation
Digital tools transform process validation in 2026 across development and manufacturing. Data historians collect continuous parameters and support faster investigations and reviews. Moreover, advanced analytics flag subtle trends before deviations impact critical quality attributes. Therefore, teams integrate sensors, MES, and LIMS into one real-time monitoring ecosystem.
Modern systems also streamline validation documentation pharma requirements globally. Structured templates, e-signatures, and audit trails strengthen data integrity and traceability. Additionally, cloud platforms centralize protocols, reports, and CPV dashboards for global sites.
Process Validation Approaches in EU GMP
Annex 15 recognises several ways to validate a process. In the traditional approach, a number of batches are made under routine conditions to confirm reproducibility. Each manufacturer must justify the number of batches using quality risk management, although a minimum of three consecutive batches is generally considered acceptable, and these may need to be supplemented with later data from ongoing process verification. For site transfers, bracketing can reduce the number of validation batches, provided existing product knowledge and the previous validation are available.
For products developed through quality by design, continuous process verification can replace traditional validation when development has shown that the control strategy gives a high degree of assurance. In that case, Process Analytical Technology and multivariate statistical process control may be used as tools. A hybrid approach combines both when substantial manufacturing experience and historical data exist. Concurrent validation, by contrast, is allowed only in exceptional cases with a strong benefit-risk ratio for patients, and it must be justified in the validation master plan. Retrospective validation is no longer acceptable.
Process Validation Documentation
Documentation is what inspectors review first. A validation master plan (VMP) should define the site’s validation system, including the validation policy, roles and responsibilities, the status of facilities and processes, change control and deviation management, and the overall strategy, including requalification.
Each process validation protocol should define the critical process parameters, critical quality attributes and their acceptance criteria, based on development data. It should also cover the equipment and its calibration status, analytical methods, in-process controls, the sampling plan and its rationale, and how results will be recorded and evaluated. Significant changes during execution, and any result that fails the acceptance criteria, must be recorded as deviations and investigated.
Finally, the validation report summarises results against the acceptance criteria and makes a clear recommendation, and responsible personnel then formally authorise release to the next stage. After that, ongoing process verification runs under its own protocol and report, and its conclusions support the validated status in the Product Quality Review.
Maintaining Process Validation
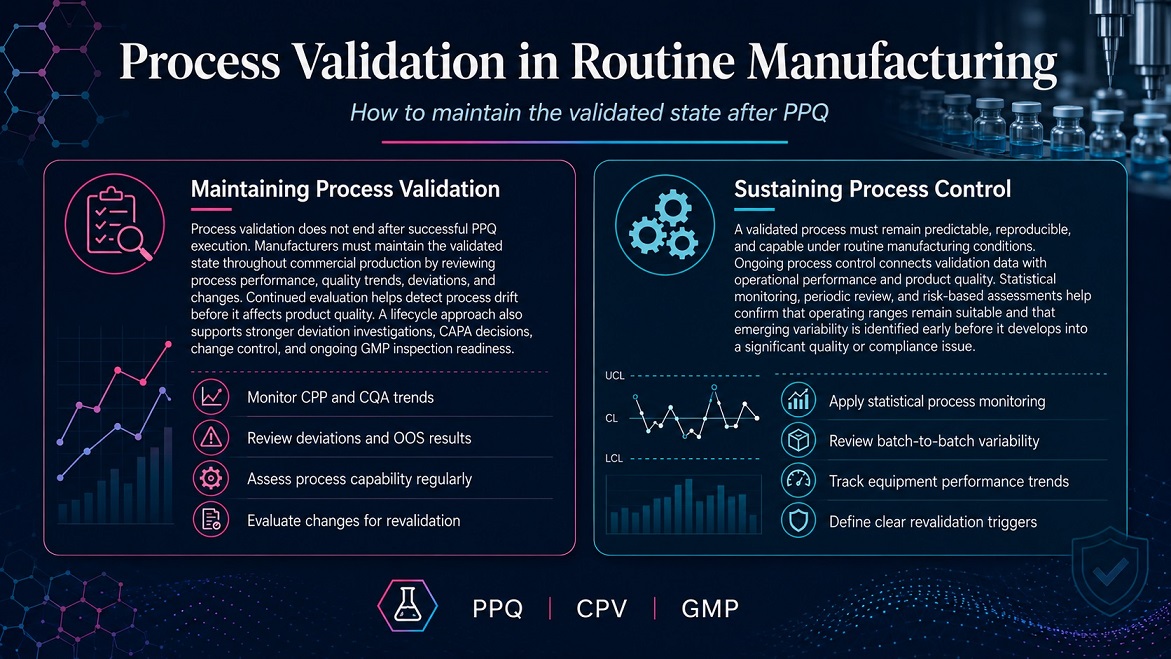
Process validation does not end after successful PPQ execution. Manufacturers must maintain the validated state throughout commercial production by reviewing process performance, quality trends, deviations, and changes. Continued evaluation helps detect process drift before it affects product quality. A lifecycle approach also supports stronger deviation investigations, CAPA decisions, change control, and ongoing GMP inspection readiness.
- Monitor CPP and CQA trends
- Review deviations and OOS results
- Assess process capability regularly
- Evaluate changes for revalidation
- Apply statistical process monitoring
- Review batch-to-batch variability
- Track equipment performance trends
- Define clear revalidation triggers
Master Process Validation Stages
Final words
Process Validation in Pharmaceutical links science, risk, and data across the whole lifecycle. It follows FDA 2011 guidance and ICH Q8, Q9, and Q10. Moreover, it uses a validation master plan to align goals, scope, and responsibilities.
Teams write each validation protocol to define tests, batches, and acceptance criteria. They collect data from Stage 1 to Stage 3 and review trends. Therefore, they support safe, consistent products for millions of patients worldwide.
To grow skills, professionals explore pharma courses on validation and GMP. They study lifecycle case studies, modern digital tools, and global inspection expectations.
FAQs
1️⃣ What are the three stages of process validation?
- FDA describes three stages. Stage 1, process design, defines the commercial process from development knowledge.
- Stage 2, process qualification, confirms that the design can reproduce quality product at commercial scale through qualified equipment and PPQ batches.
- Stage 3, continued process verification, provides ongoing assurance during routine production that the process stays in a state of control.
2️⃣ Is concurrent validation allowed?
Only in exceptional circumstances with a strong benefit-risk ratio for patients.
Under EU GMP Annex 15, the decision must be justified, documented in the validation master plan and approved, and there must be enough data to show each batch meets the acceptance criteria before the Qualified Person certifies it.
3️⃣ How do Stage 2 and Stage 3 support routine manufacturing?
Stage 2 PPQ shows the process runs consistently at commercial scale.
Stage 3 CPV tracks trends, deviations, and improvements over time.
References
Ershad Moradi, a Content Marketing Specialist at Zamann Pharma Support, brings 6 years of experience in the pharmaceutical industry. Specializing in pharmaceutical and medical technologies, Ershad is currently focused on expanding his knowledge in marketing and improving communication in the field. Outside of work, Ershad enjoys reading and attending industry related networks to stay up-to-date on the latest advancements. With a passion for continuous learning and growth, Ershad is always looking for new opportunities to enhance his skills and contribute to pharmaceutical industry. Connect with Ershad on Facebook for more information.
Pharmaceutical Cold Chain Excursion: Causes, Impact, and a Practical Management Guide
A pharmaceutical cold chain excursion can threaten medicine quality, stability, and patient safety. Learn how to detect, document, assess, and manage temperature deviations using current guidance and science-based risk assessment.
Pharmaceutical Supply Chain: The Ultimate Guide to Hidden Risks and Challenges
Protocol deviations can threaten participant protection and trial reliability. This practical guide explains risk-based classification, documentation, escalation, CAPA, and preventive oversight.
Protocol Deviations in Clinical Trials: Build an Audit-Ready Record
Protocol deviations can threaten participant protection and trial reliability. This practical guide explains risk-based classification, documentation, escalation, CAPA, and preventive oversight.