


FDA good manufacturing practices are the current good manufacturing practice (CGMP) rules that set minimum requirements for the methods, facilities and controls used to make medicines. For drugs, they sit in 21 CFR Parts 210 and 211, while medical devices follow a separate framework under Part 820.
Good Manufacturing Practices turn these principles into controls that an inspector can verify. In FY2023, FDA added 93 companies to import alerts for drug-quality violations, compared with 77 across FY2021 and FY2022 combined. FDA also reported 674 recalled drug products in FY2023, down 26% from 912 in FY2022. However, these historical figures describe enforcement and recall activity; they do not show that stronger quality systems caused the decline.
Table of Contents
Inside the FDA Good Manufacturing Practices Framework
In daily work, FDA good manufacturing practices mean following approved procedures, controlling changes and keeping records that show what actually happened. The aim is to prevent contamination, mix-ups and errors before they affect a batch.
Drug GMP regulations appear in 21 CFR Parts 210 and 211. These parts require validated processes, controlled facilities, and qualified personnel. They govern documentation, labeling, testing, and stability. For devices, 21 CFR Part 820 sets the Quality System Regulation. The rules firmly anchor FDA compliance across the lifecycle.
GMP guidelines set the foundation for quality operations. Companies train people, validate methods, and keep complete records. Digital QMS platforms centralize SOPs, CAPA, and change control. Teams run risk-based audits and qualify suppliers. Additionally, continuous improvement keeps systems current with cGMP expectations and strengthens inspection readiness and compliance.
Historical Background of FDA GMP
FDA introduced GMP after early safety concerns and legislative reforms. In 1963, FDA published the first Good Manufacturing Practice regulations for drugs. They later codified cGMP within 21 CFR Parts 210 and 211. For devices, FDA established Part 820 in 1978 and revised it in 1996. These milestones made GMP enforceable across manufacturing. Moreover, regulators tied approvals and inspections to robust, documented quality controls.
Therefore, GMP became mandatory to protect patients and assure consistent, quality outcomes.
Prevent contamination, mix-ups, and labeling errors.
Prove processes work through validation and documented controls.
Qualify people, facilities, equipment, and suppliers.
Enable traceability, recalls, and sustained FDA compliance.
See 21 CFR Parts 210, 211, and 820 for scope.
The Meaning of "Current" in FDA CGMP
The “C” in CGMP stands for current. FDA expects manufacturers to use up-to-date technologies and systems, so equipment that was adequate ten or twenty years ago may no longer meet today’s standard.
The table below compares three common beliefs with what the CGMP regulations actually say.
| Common belief | What FDA CGMP actually says |
|---|---|
|
CGMP is a stricter tier above GMP |
“Current” describes how GMP is applied; GMP and CGMP are not separate regulatory tiers |
|
Every site needs electronic batch records or a digital QMS |
The regulations do not require a specific technology; they require effective controls and complete records |
|
The rules prescribe exactly how to manufacture |
The regulations set minimum requirements and let each manufacturer decide how to implement them using sound science |
FDA GMP Requirements for Pharmaceuticals and Medical Devices
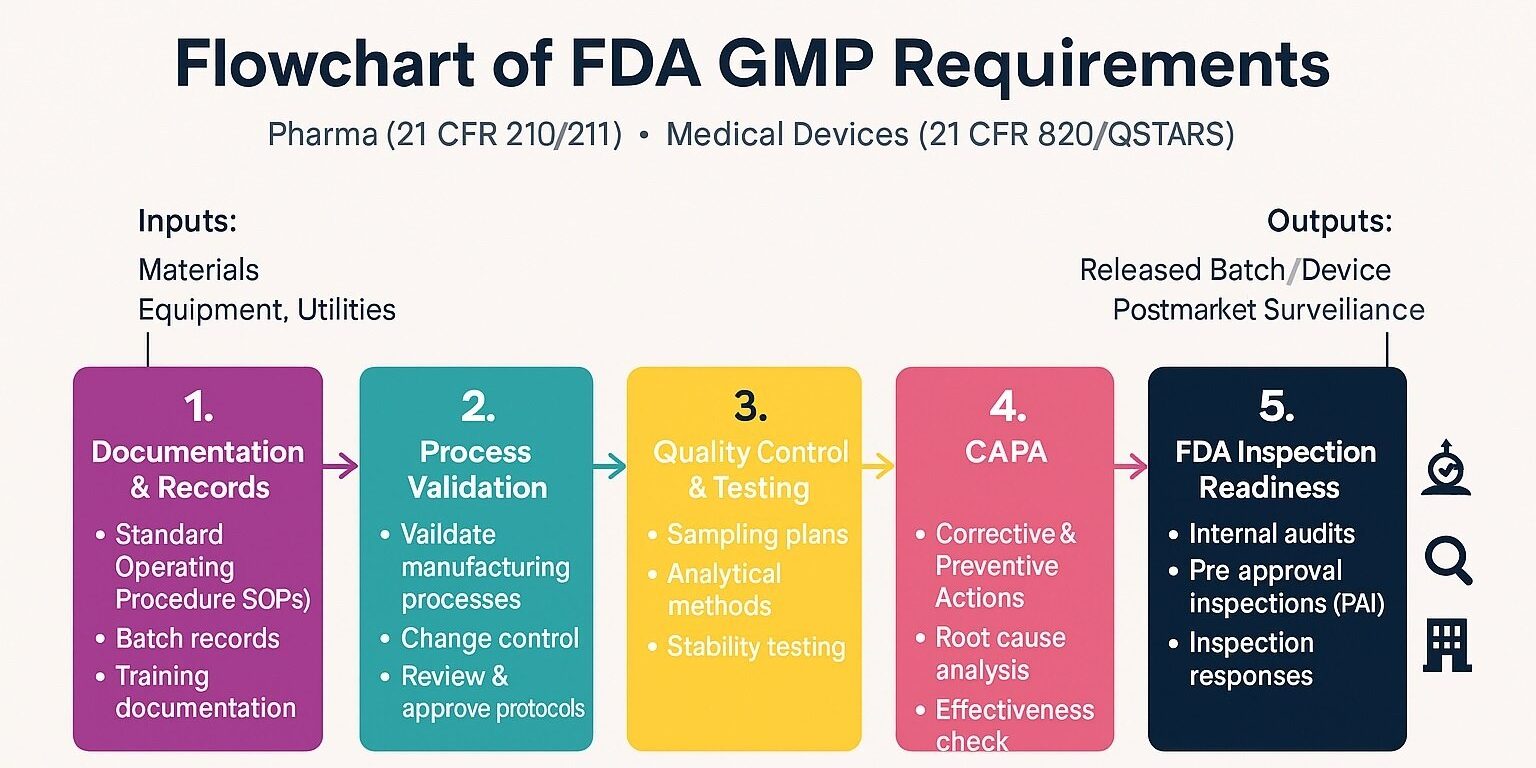
FDA GMP requirements depend on the product type. Drugs follow 21 CFR Parts 210 and 211, while devices follow 21 CFR Part 820, now the Quality Management System Regulation (QMSR). For drugs, manufacturers document their work, validate processes and maintain traceability, and an independent quality unit reviews the records before product is released. Teams also manage risk through CAPA, change control and trending, qualify suppliers against specifications, and validate electronic systems to protect data integrity. Training and inspection readiness run through all of this rather than sitting in a separate project.
For drug manufacturers, four areas shape most of the daily work:
Documentation and training: write SOPs, maintain records, and verify competency.
Process validation: prove processes and cleaning deliver consistent, compliant outcomes.
Hygiene and facilities: control environments, utilities, and equipment to prevent contamination.
Quality control testing: sample, test, and authorize batches before release.
How to Ensure Compliance with FDA GMP
Internal Audits
Schedule routine self-inspections across production, labs, warehouses, and quality units.
Staff Training
Map competencies to roles. Train on SOPs, data integrity principles, and GMP regulations.
Supplier Qualification
Risk-rank suppliers. Perform paper or on-site audits. Approve only those who meet defined criteria.
CAPA System
Establish a structured Corrective and Preventive Action process. Investigate root causes using sound methods.
Mock FDA Inspections
Conduct realistic simulations with trained auditors. Practice document retrieval, interview skills, and tour sequences.
Finally, choose record-management tools that fit the site’s operations and the requirements that apply to it. A digital QMS can help organize SOPs, deviations, CAPA and audit findings, but buying software does not, on its own, establish CGMP compliance. Whichever system you use, records must stay readily available for FDA inspection throughout their retention period, as 21 CFR 211.180(c) requires.
FDA GMP Inspection Readiness Checklist
FDA inspections focus on evidence, not only written procedures. Before an inspection, companies should confirm that key records are current, complete, approved, and easy to retrieve. The review should cover production, quality control, validation, training, equipment, suppliers, deviations, CAPA, complaints, and data integrity. A structured readiness check helps teams identify missing evidence, assign corrective actions, and reduce delays during document requests. It also helps employees explain how daily controls support product quality and patient safety across all affected departments consistently.
- Confirm that SOPs, batch records, and laboratory data are complete and approved.
- Review deviations, CAPA actions, validation reports, and training records.
- Test whether requested documents can be retrieved quickly during an inspection.
21 CFR 210 and 211: From Regulation to Daily Controls
21 CFR Parts 210 and 211 define the core FDA requirements for manufacturing finished pharmaceutical products. Part 210 explains the general scope and applicability of current good manufacturing practice, while Part 211 provides detailed requirements for personnel, facilities, equipment, production, laboratories, records, labeling, and complaints. Companies should translate these regulations into controlled procedures, assigned responsibilities, monitoring, and documented evidence. This practical approach helps employees understand not only what the regulations require, but how compliance is demonstrated during operations and inspections.
- Connect each regulatory requirement with a controlled procedure and responsible department.
- Maintain documented evidence for production, laboratory, equipment, and quality activities.
- Review controls regularly to confirm that they remain effective and compliant.

A Passing Test Does Not Establish CGMP Compliance
A finished-product result speaks only for the samples tested. It cannot show that every manufacturing step was controlled, because FDA expects quality to be built into the process rather than tested into the product. A batch that meets its specification can therefore still raise CGMP concerns.
Imagine a batch that passes release testing despite an unexplained interruption in its production record. Rather than testing another sample, find out what happened, which controls were affected and whether the records support release. Under 21 CFR 211.192, the quality control unit reviews these records before release, and unexplained discrepancies must be investigated even after distribution.
Connect a Discrepancy to the Wider Product Review
An investigation should not stop at the first batch. Under 21 CFR 211.192, it extends to other batches and products that may be associated with the discrepancy, so list shared equipment, materials or steps and record which batches were included and why.
The findings then feed the annual product review. 21 CFR 211.180(e) requires at least a yearly evaluation that brings batches, complaints, recalls and investigations together, which can reveal a recurring issue that single records hide.
Get Your 21 CFR 210 and 211 Checklist
Final Words
FDA good manufacturing practices ultimately come down to effective controls backed by complete records and justified quality decisions. At the end of FY2023, CDER’s Site Catalog contained 4,819 drug-manufacturing sites, and among those with an inspection outcome, 94% had a most recent classification of No Action Indicated or Voluntary Action Indicated. These are historical site-level classifications, not a product pass rate. For medical devices, meanwhile, the QMSR has applied since February 2, 2026, bringing Part 820 into line with ISO 13485.
FAQs
1️⃣ What are FDA GMP and cGMP?
Answer: GMP and CGMP refer to the same FDA rules, which set requirements for the methods, facilities and controls that support a drug’s identity, strength, quality and purity. “Current” means using up-to-date methods and systems. However, CGMP compliance alone does not establish a drug’s clinical effectiveness.
2️⃣ Who must follow them, and where are they written?
Answer: Drug manufacturers follow 21 CFR Parts 210–211; medical-device manufacturers follow 21 CFR Part 820 (now the Quality Management System Regulation, QMSR). These regulations apply across the lifecycle materials, production, packaging/labeling, testing, release, distribution, and complaints for both U.S. and foreign facilities that supply the U.S. market.
3️⃣ How do we show compliance in practice?
Answer: Build a documented, auditable quality system that includes:
Written SOPs, complete batch/device records, and data integrity controls.
Validated processes, methods, cleaning, and computerized systems.
Qualified personnel, role-based training, and independent quality control. Investigations with documented conclusions and follow-up, supported by complete production and control records.
References
Ershad Moradi, a Content Marketing Specialist at Zamann Pharma Support, brings 6 years of experience in the pharmaceutical industry. Specializing in pharmaceutical and medical technologies, Ershad is currently focused on expanding his knowledge in marketing and improving communication in the field. Outside of work, Ershad enjoys reading and attending industry related networks to stay up-to-date on the latest advancements. With a passion for continuous learning and growth, Ershad is always looking for new opportunities to enhance his skills and contribute to pharmaceutical industry. Connect with Ershad on Facebook for more information.
eCTD Publishing Software: The Right Features for Submission Success
Discover how eCTD publishing software organizes regulatory dossiers, validates electronic submissions, manages document lifecycles, and supports FDA and EMA workflows.
Pharmaceutical Analytical Testing Services: Essential Quality Insights
A practical guide to pharmaceutical analytical testing services: assay, impurities, dissolution, stability, microbial testing, validation and outsourced-laboratory oversight.
Medical Device Distribution Quality: Essential Safeguards Against Failures
Medical device distribution quality protects device conformity from receipt to delivery. Explore EU distributor obligations, ISO 13485 principles, UDI traceability, storage controls, complaint handling, and audit-ready records.